Vol. 13. Núm. 79.
Páginas 4685-4694 (Mayo 2023)

Actualización
Afectación renal en las enfermedades raras genéticas

Renal involvement in rare genetic diseases

P. Rodríguez DoyágüezaN.B. BoldobaaR. GimenabB. Hernández SevillanoaR. Torra Ballcellsc
Doi : https://10.1016/j.med.2023.05.004
Opciones
Resumen

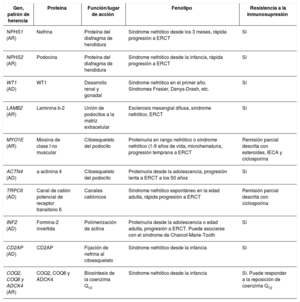
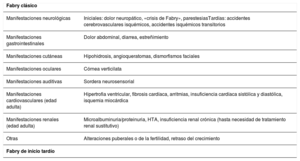
Las enfermedades raras se definen como aquellas que afectan a menos de 65 de cada 100 000 personas. Las enfermedades genéticas que afectan al riñón son, en su mayoría, pertenecientes a ese grupo y constituyen un porcentaje significativo de los pacientes en tratamiento renal sustitutivo, especialmente a edades tempranas. Junto con las enfermedades quísticas, las tubulopatías o las mutaciones en genes del complemento con afectación renal existen enfermedades renales genéticas con afectación a distintos niveles. Algunas de ellas tienen como diana el glomérulo, como el síndrome de Alport (que afecta a las proteínas que forman el colágeno tipo IV, cursando con proteinuria, microhematuria y alteraciones visuales y auditivas), la amiloidosis hereditaria (que cursa como la amiloidosis AL y es responsable del 10% total de casos) y el síndrome nefrótico corticorresistente que surge de diversas mutaciones y es resistente a la inmunosupresión. La enfermedad de Fabry provoca la acumulación de glucoesfingolípidos, produciendo una afectación multisistémica con mortalidad cardíaca y renal. La hiperoxaluria primaria destaca por urolitiasis de repetición y nefrocalcinosis con afectación multisistémica cuando hay insuficiencia renal avanzada. Finalmente, el CAKUT comprende un conjunto heterogéneo de mutaciones de genes responsables del desarrollo embrionario del riñón y el tracto urinario.

Palabras clave
Genética
Monogénica
Síndrome de Alport
Abstract

Rare diseases are defined as those which affect fewer than 65 of every 100,000 individuals. Genetic diseases which affect the kidney mostly belong to this group and represent a significant percentage of patients in renal replacement treatment, especially at young ages. Together with cystic diseases, tubulopathies, or complement gene mutations with renal involvement, there are genetic renal diseases which affect different aspects. Some of them target the glomerulus, such as Alport syndrome (which affects the proteins that form type IV collagen, progressing with proteinuria, microscopic hematuria, and visual and auditory abnormalities), hereditary amyloidosis (which progress with AL amyloidosis and is responsible for 10% of all cases), and steroid-resistant nephrotic syndrome which arises from various mutations and is resistant to immunosuppression. Fabry disease causes the accumulation of glycosphingolipids, producing multisystemic involvement with cardiac and renal mortality. Primary hyperoxaluria is notable due to recurrent urolithiasis and nephrocalcinosis with multisystemic involvement when there is end-stage renal failure. Finally, CAKUT comprises a heterogeneous set of mutations in the genes responsible for the embryonic development of the kidney and urinary tract.

Keywords
Genetics
Monogenetic
Alport syndrome

Identifíquese

¿Aún no es suscriptor de la revista?
Comprar el acceso al artículo

Comprando el artículo el pdf del mismo podrá ser descargado

Teléfono para incidencias

De lunes a viernes de 9h a 18h (GMT+1) excepto los meses de julio y agosto que será de 9 a 15h
Llamadas desde España 932 415 960
Llamadas desde fuera de España +34 932 415 960
Bibliografía
[1]
M.F.M.D. Cunha
202305231130179858
, G. Sevignani, G.M. Pavanelli, M. de Carvalho, F.C. Barreto.
Rare inherited kidney diseases: an evolving field in Nephrology.
J Bras Nefrol., (2020), 42 pp. 219-230
[2]
E. Ars
202305231130179859
, R. Torra.
Rare diseases, rare presentations: recognizing atypical inherited kidney disease phenotypes in the age of genomics.
Clin Kidney J., (2017), 10 pp. 586-593
[3]
N. Knoers
2023052311301798510
, C. Antignac, C. Bergmann, K. Dahan, S. Giglio, L. Heidet, et al.
Genetic testing in the diagnosis of chronic kidney disease: recommendations for clinical practice.
Nephrol Dial Transplant., (2022), 37 pp. 239-254
[4]
B.Q. Gili
2023052311301798511
, K.M.P. del Valle, B.H. Sevillano, G. de la Fuente.
Nefropatías congénitas y hereditarias (I): tubulopatías.
Medicine., (2019), 12 pp. 4878-4884
[5]
K.M.P. del Valle
2023052311301798512
, B.H. Sevillano, D.G.G. Tocora, B.Q. Gili, G.de A. de la Fuente.
Nefropatías congénitas y hereditarias (II): enfermedad quística.
Medicine [Internet]., (2019), 12 pp. 4885-4894
[6]
M. Furlano
2023052311301798513
, V. Martínez, M. Pybus, Y. Arce, J. Crespí, M.D.P. Venegas, et al.
Clinical and genetic features of autosomal dominant Alport syndrome: A cohort study.
Am J Kidney Dis [Internet]., (2021), 78 pp. 560-5700
[7]
J. Savige
2023052311301798514
, F. Ariani, F. Mari, M. Bruttini, A. Renieri, O. Gross, et al.
Expert consensus guidelines for the genetic diagnosis of Alport syndrome.
Pediatr Nephrol., (2019), 34 pp. 1175-1189
[8]
J.M. Lee
2023052311301798515
, K. Nozu, D.E. Choi, H.G. Kang, I.-S. Ha, H. Cheong.
Features of autosomal recessive Alport syndrome: A systematic review.
J Clin Med [Internet]., (2019), 8 pp. 178
[9]
J. Savige
2023052311301798516
, M. Gregory, O. Gross, C. Kashtan, J. Ding, F. Flinter.
Expert guidelines for the management of Alport syndrome and thin basement membrane nephropathy.
J Am Soc Nephrol [Internet]., (2013), 24 pp. 364-375
[10]
C. Fallerini
2023052311301798517
, M. Baldassarri, E. Trevisson, V. Morbidoni, A. La Manna, R. Lazzarin, et al.
Alport syndrome: impact of digenic inheritance in patients management.
Clin Genet [Internet]., (2017), 92 pp. 34-44
[11]
L. Heidet
2023052311301798518
, M.-C. Gubler.
The renal lesions of Alport syndrome.
J Am Soc Nephrol [Internet]., (2009), 20 pp. 1210-1215
[12]
M. Furlano
2023052311301798519
, R. Arlandis, M.D.P. Venegas, S. Novelli, J. Crespi, G. Bullich, et al.
Nefropatía asociada a mutación del gen MYH9.
Nefrol (Engl Ed) [Internet]., (2019), 39 pp. 133-140
[13]
I.D. Liu
2023052311301798520
, N.S. Willis, J.C. Craig, E.M. Hodson.
Interventions for idiopathic steroid-resistant nephrotic syndrome in children.
Cochrane Database Syst Rev [Internet]., (2019), 2019
[14]
J.M. Lee
2023052311301798521
, A. Kronbichler, J.I. Shin, J. Oh.
Current understandings in treating children with steroid-resistant nephrotic syndrome.
Pediatr Nephrol [Internet]., (2021), 36 pp. 747-761
[15]
J.K. Warejko
2023052311301798522
, W. Tan, A. Daga, D. Schapiro, J.A. Lawson, S. Shril, et al.
Whole exome sequencing of patients with steroid-resistant nephrotic syndrome.
Clin J Am Soc Nephrol [Internet]., (2018), 13 pp. 53-62
[16]
J.B. Kopp
2023052311301798523
, H.-J. Anders, K. Susztak, M.A. Podestà, G. Remuzzi, F. Hildebrandt, et al.
Podocytopathies.
Nat Rev Dis Primers [Internet], (2020), 6 pp. 68
[17]
O. Boyer
2023052311301798524
, G. Dorval, A. Servais.
Hereditary podocytopathies in adults: The next generation.
Kidney Dis (Basel) [Internet]., (2017), 3 pp. 50-56
[18]
L.A. Bruggeman
2023052311301798525
, J.F. O’Toole, J.R. Sedor.
APOL1 polymorphisms and kidney disease: loss-of-function or gain-of-function?.
Am J Physiol Renal Physiol [Internet]., (2019), 316 pp. F1-F8
[19]
Y. Harita
2023052311301798526
, S. Urae, R. Akashio, T. Isojima, K. Miura, T. Yamada, et al.
Clinical and genetic characterization of nephropathy in patients with nail-patella syndrome.
Eur J Hum Genet [Internet]., (2020), 28 pp. 1414-1421
[20]
M.M. Picken
2023052311301798527
.
The pathology of amyloidosis in classification: A review.
Acta Haematol [Internet]., (2020), 143 pp. 322-334
[21]
R. Ryšavá
2023052311301798528
.
AL amyloidosis: advances in diagnostics and treatment.
Nephrol Dial Transplant [Internet]., (2019), 34 pp. 1460-1466
[22]
A. Felis
2023052311301798529
, M. Whitlow, A. Kraus, D.G. Warnock, E. Wallace.
Current and investigational therapeutics for Fabry disease.
Kidney Int Rep [Internet]., (2020), 5 pp. 407-413
[23]
E. Viggiano
2023052311301798530
, L. Politano.
X chromosome inactivation in carriers of Fabry disease: Review and meta-analysis.
Int J Mol Sci [Internet]., (2021), 22 pp. 7663
[24]
M. Lenders
2023052311301798531
, E. Brand.
Precision medicine in Fabry disease.
Nephrol Dial Transplant [Internet]., (2021), 36 pp. 14-23
[25]
A. Ortiz
2023052311301798532
, D.P. Germain, R.J. Desnick, J. Politei, M. Mauer, A. Burlina, et al.
Fabry disease revisited: Management and treatment recommendations for adult patients.
Mol Genet Metab [Internet]., (2018), 123 pp. 416-427
[26]
M. Řeboun
2023052311301798533
, J. Sikora, M. Magner, H. Wiederlechnerová, A. Černá, H. Poup¿tová, et al.
Pitfalls of X-chromosome inactivation testing in females with Fabry disease.
Am J Med Genet A [Internet]., (2022), 188 pp. 1979-1989
[27]
D.S. Milliner
2023052311301798534
, P.C. Harris, D.J. Sas, A.G. Cogal, J.C. Lieske.
Primary hyperoxaluria type 1.
GeneReviews® [Internet], University of Washington, Seattle; 1993-2022,
[28]
G. Rumsby
2023052311301798535
, S.A. Hulton.
Primary hyperoxaluria type 2.
GeneReviews® [Internet], University of Washington, Seattle; 1993-2022,
[29]
E.L. Metry, L.M.M. van Dijk, H. Peters-Sengers, M.J.S. Oosterveld, J.W. Groothoff, R.J. Ploeg, et al.
Transplantation outcomes in patients with primary hyperoxaluria: a systematic review.
Pediatr Nephrol [Internet]., (2021), 36 pp. 2217-2226
[30]
S.F. Garrelfs
2023052311301798536
, Y. Frishberg, S.A. Hulton, M.J. Koren, W.D. O’Riordan, P. Cochat, et al.
Lumasiran, an RNAi therapeutic for primary hyperoxaluria type 1.
N Engl J Med [Internet]., (2021), 384 pp. 1216-1226
[31]
A.T. van der Ven
2023052311301798537
, D.M. Connaughton, H. Ityel, N. Mann, M. Nakayama, J. Chen, et al.
Whole-exome sequencing identifies causative mutations in families with congenital anomalies of the kidney and urinary tract.
J Am Soc Nephrol [Internet]., (2018), 29 pp. 2348-2361
[32]
N.V.A.M. Knoers
2023052311301798538
, K.Y. Renkema.
The genomic landscape of CAKUT; you gain some, you lose some.
Kidney Int [Internet]., (2019), 96 pp. 267-269
[33]
R.L. Clissold
2023052311301798539
, A.J. Hamilton, A.T. Hattersley, S. Ellard, C. Bingham.
HNF1B-associated renal and extra-renal disease—an expanding clinical spectrum.
Nat Rev Nephrol [Internet]., (2015), 11 pp. 102-112